danhumassmed/qc-tools
Software for Bioinformatics pipelines FastQC, MultiQC, RSeQC
256
danhumassmed/qc-tools repository overview
Sofware Provided
| Docker Tag | FastQC | MultiQC | RSeQC |
|---|---|---|---|
| 1.0.1 | v0.12.1 | version 1.14 | 4.0.0 |
| 1.0.2 | v0.12.1 | version 1.26 | 4.0.0 |
FastQC
https://www.bioinformatics.babraham.ac.uk/projects/fastqc/
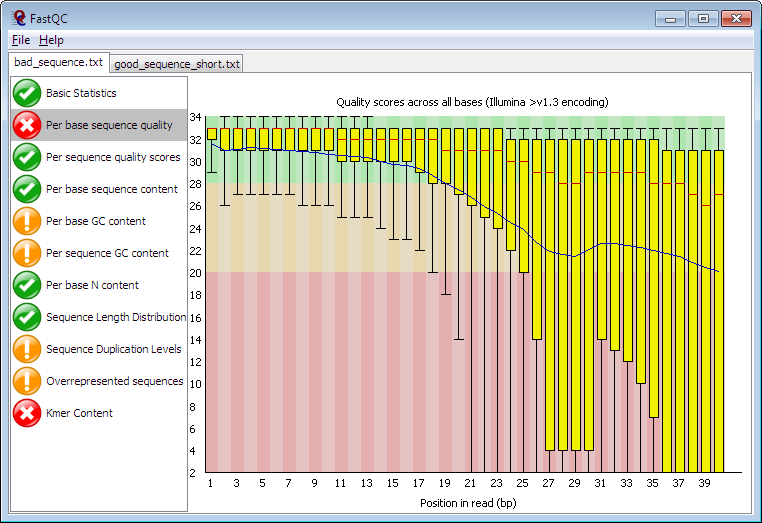
FastQC aims to provide a simple way to do some quality control checks on raw sequence data coming from high throughput sequencing pipelines. It provides a modular set of analyses which you can use to give a quick impression of whether your data has any problems of which you should be aware before doing any further analysis.
The main functions of FastQC are:
- Import of data from BAM, SAM or FastQ files (any variant)
- Providing a quick overview to tell you in which areas there may be problems
- Summary graphs and tables to quickly assess your data
- Export of results to an HTML based permanent report
- Offline operation to allow automated generation of reports without running the interactive application
MultiQC
Aggregate results from bioinformatics analyses across many samples into a single report MultiQC searches a given directory for analysis logs and compiles a HTML report. It's a general use tool, perfect for summarising the output from numerous bioinformatics tools.

RSeQC
https://pythonhosted.org/RSeQC//

RSeQC package provides a number of useful modules that can comprehensively evaluate high throughput sequence data especially RNA-seq data. Some basic modules quickly inspect sequence quality, nucleotide composition bias, PCR bias and GC bias, while RNA-seq specific modules evaluate sequencing saturation, mapped reads distribution, coverage uniformity, strand specificity, transcript level RNA integrity etc.
Usage
The provided Docker image is compatible with Singularity and is actively used in NextFlow Pipelines configured for an HPC.
Example running fastqc (Mac OSX)
local_project_dir="/Users/dan/Code/NextFlow/Bioinformatics_Training/Introduction_to_RNA-seq"
docker_project_dir="/home/rnaseq"
input_dir="raw_data"
output_dir="results/fastqc/"

mkdir -p ${local_project_dir}/${output_dir}
docker run -v ${local_project_dir}:${docker_project_dir} danhumassmed/qc-tools:1.0.1 \
    /bin/bash -c \"fastqc -o ${docker_project_dir}/${output_dir} -f fastq ${docker_project_dir}/${input_dir}/*.fq\"
Tag summary
Content type
Image
Digest
sha256:9a3bf7986…
Size
1002.2 MB
Last updated
over 1 year ago
docker pull danhumassmed/qc-tools:1.0.2